https://lmspiq.fda.gov.tw/web/MDPIQ/MDPIQ1000Result?licBaseId=7062758E-564E-4EFF-BD9F-EBB74A98192F
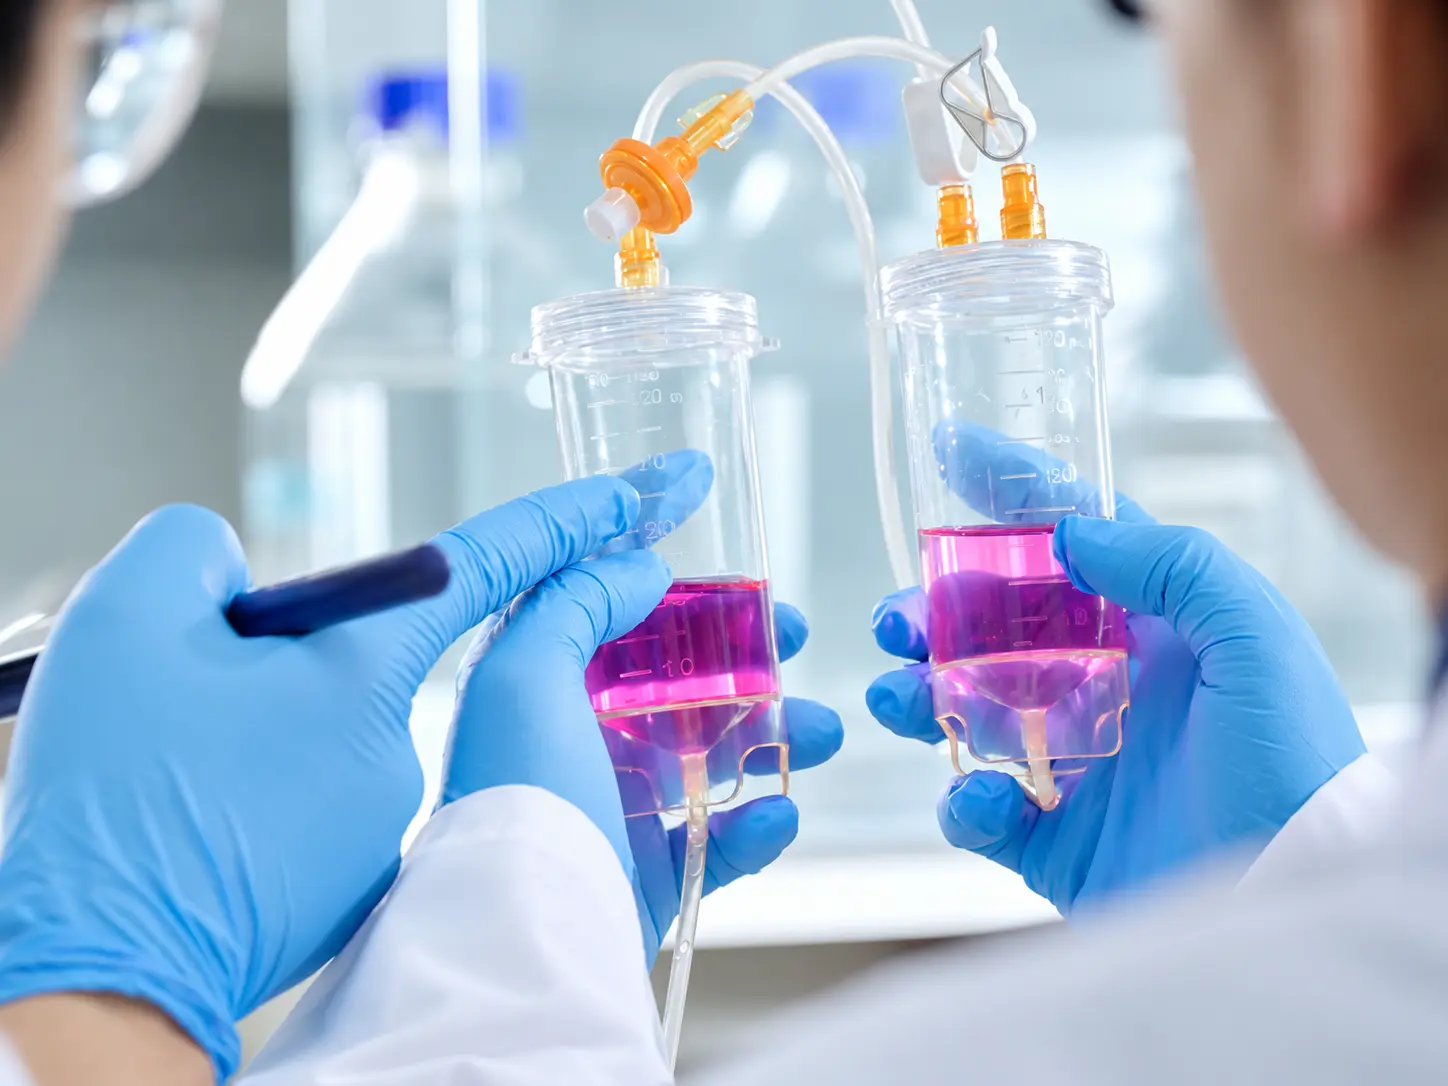
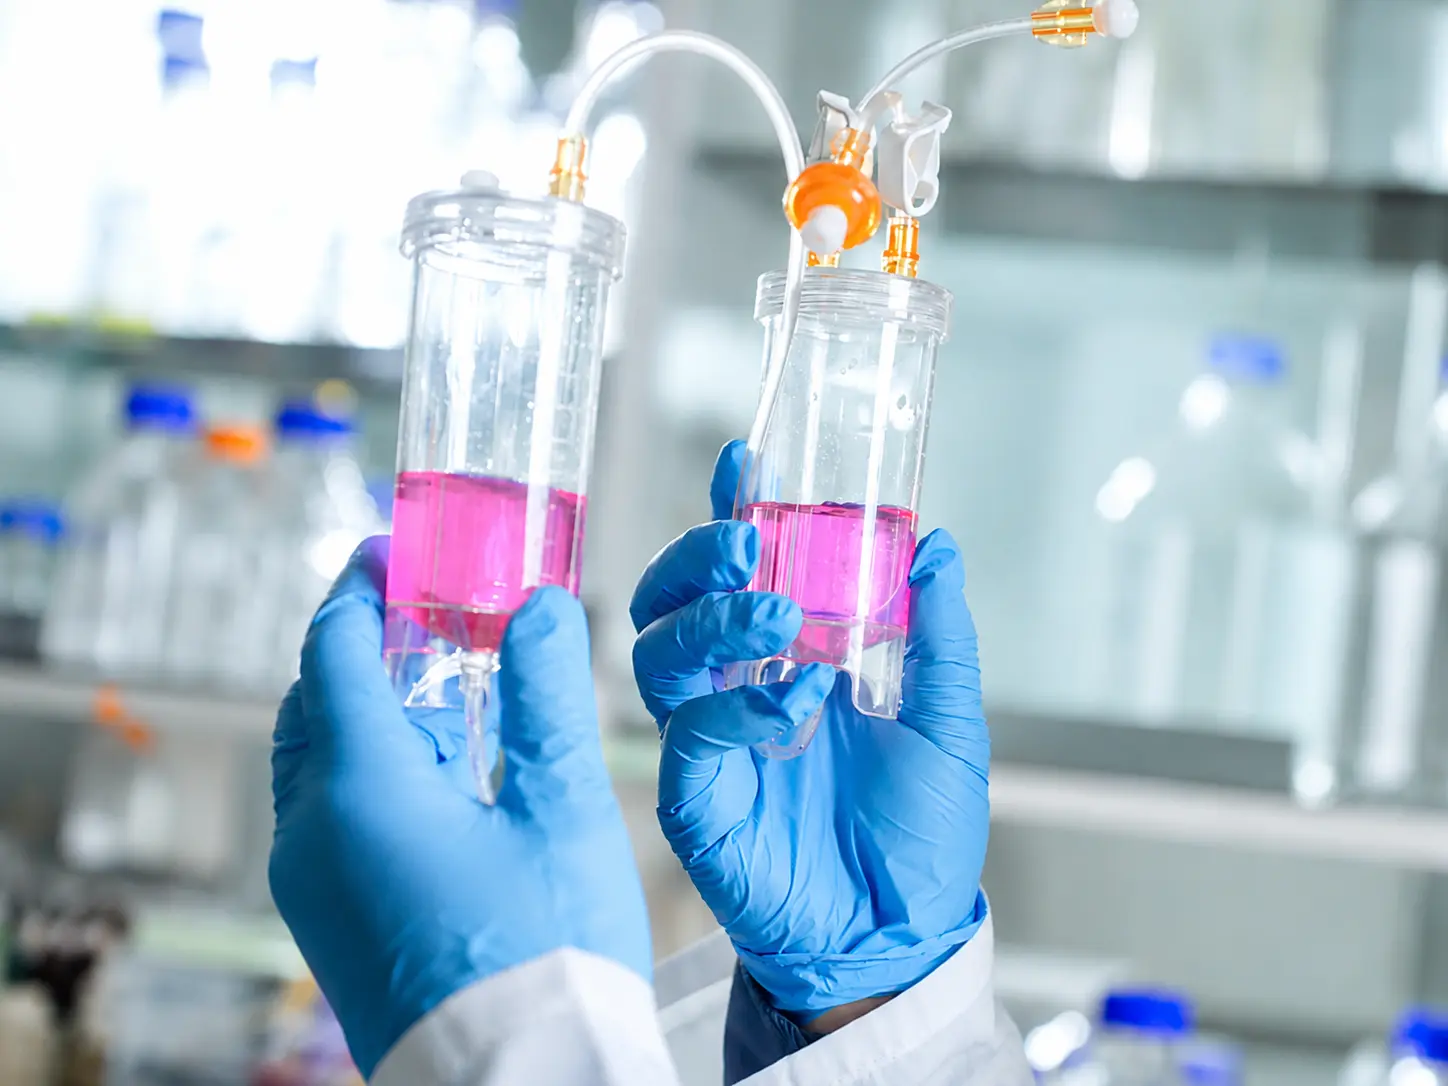
"Nanchac" Blood Components Separation Kit (007629)
USA FDA 510(k) number (K240247)
DuoGenic StemCells corporation
QMS1872
GMP ISO13485
The product underwent verification and validation throughout development and manufacturing to ensure safety, performance, and quality consistency. Third-party test results are summarized below: 1. Biocompatibility (ISO 10993): Cytotoxicity, irritation, sensitization, hemolysis, pyrogenicity, acute systemic toxicity, and endotoxin tests showed no biocompatibility concerns. 2. Hemocompatibility (ISO 10993-4): Coagulation, platelet, and hematological analyses confirmed no coagulation abnormalities. 3. Particulate and Physicochemical Tests: Particulates, reducing substances, ammonia, chloride, heavy metals, pH, residues, UV absorbance, and plasticizers met blood centrifuge tube requirements. DEHP was not detected (ISO 3826-4:2015). Based on worst-case assessment, use is not recommended for individuals weighing <21 kg. 4. Sterilization Validation: The sterilization process achieved a Sterility Assurance Level (SAL) of 10⁻⁶. 5. Real-Time Aging (18 months): Seal strength, dye penetration, burst, microbial barrier, and sterility tests confirmed maintenance of sterile barrier integrity. 6. Microbial Barrier Test: Effective microbial barrier performance was demonstrated using Brevundimonas diminuta. 7. Functional Evaluation: Destructive centrifugation, centrifugal force, and centrifuge compatibility tests established the recommended operating conditions and compatible centrifuge models.
ISO 13485: 2016
ISO 9001:2015
GMP/QMS
Raw Material Control and Inspection
In-Process Inspection
Finished Product Inspection
Safety and Efficacy Testing
Stability Testing
Post-Market Surveillance
The post-market surveillance system covers safety monitoring, crisis handling, quality improvement, and CAPA to ensure continued safety and performance after commercialization.
Product Monitoring and Feedback: Market surveys, clinical follow-up, and user satisfaction questionnaires are used to collect feedback during routine use, supporting early detection of design issues, usability problems, or potential risks.
Adverse Event Reporting: Any serious injury, death, or product malfunction is reported to the competent authority within the required timeframe per regulations.
Risk Re-evaluation: Post-market data are reviewed against the ISO 14971 risk management file to reassess probability and severity, ensuring clinical benefits outweigh risks and supporting decisions on modification or recall.
Advisory Notice and Recall: If safety issues are identified, advisory notices may be issued, labeling or instructions updated, and affected products corrected, withdrawn, or recalled to reduce risk.
CAPA: Root causes of adverse events or quality issues are investigated, followed by corrective actions in design, manufacturing, procurement, or quality control, and preventive measures to avoid recurrence.
Post-market Review: Relevant regulations and scientific literature are periodically reviewed to support continuous improvement and risk management.